Can an endoscope monitor display real – time images?
Hey there! As a supplier of Endoscope Monitors, I often get asked this question: Can an endoscope monitor…
Read article →
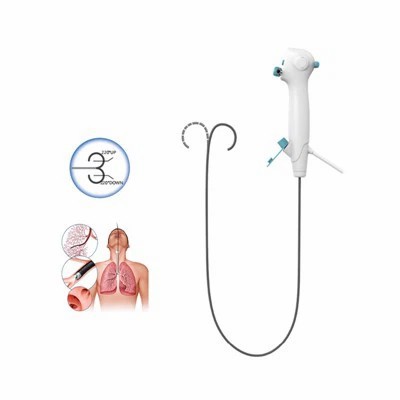
As a supplier of portable bronchoscopes, I understand the critical importance of regulatory compliance in the medical device industry. Portable bronchoscopes are essential tools in the diagnosis and treatment of various respiratory conditions, and ensuring they meet all regulatory requirements is not only a legal obligation but also a matter of patient safety and trust. In this blog post, I will delve into the key regulatory requirements that portable bronchoscopes must adhere to, providing insights into the standards and processes that govern their development, manufacturing, and distribution.
The regulatory requirements for portable bronchoscopes vary from country to country, but there are several international standards and guidelines that serve as a foundation for regulatory compliance. The International Organization for Standardization (ISO) plays a crucial role in developing standards for medical devices, including bronchoscopes. ISO 13485:2016 is an internationally recognized standard for quality management systems in the medical device industry. It provides a framework for ensuring that medical devices are designed, developed, produced, installed, and serviced in a manner that meets regulatory requirements and customer needs.
In addition to ISO standards, the World Health Organization (WHO) has also issued guidelines on the safety and performance of medical devices, including bronchoscopes. These guidelines aim to promote the availability of safe and effective medical devices worldwide, particularly in low- and middle-income countries.
In the United States, the Food and Drug Administration (FDA) is responsible for regulating medical devices, including portable bronchoscopes. The FDA classifies medical devices into three classes based on the level of risk they pose to patients: Class I (low risk), Class II (moderate risk), and Class III (high risk). Portable bronchoscopes are typically classified as Class II devices, which means they are subject to special controls in addition to general controls.
General controls include requirements for device registration, listing, labeling, and good manufacturing practices (GMP). Special controls may include performance standards, post-market surveillance, and clinical trials. To market a portable bronchoscope in the United States, manufacturers must submit a 510(k) premarket notification to the FDA, demonstrating that the device is substantially equivalent to a legally marketed device (predicate device) in terms of safety and effectiveness.
In some cases, manufacturers may be required to conduct clinical trials to support the safety and effectiveness of their portable bronchoscopes. Clinical trials involve testing the device on human subjects under controlled conditions to evaluate its performance and potential risks. The FDA reviews the results of clinical trials to determine whether the device can be approved for marketing in the United States.
In the European Union (EU), medical devices are regulated by the Medical Device Regulation (MDR) 2017/745, which came into effect on May 26, 2021. The MDR aims to improve the safety and performance of medical devices in the EU by introducing stricter regulatory requirements and enhancing transparency and traceability.
Under the MDR, portable bronchoscopes are classified as Class IIa devices, which means they are subject to conformity assessment procedures by a notified body. A notified body is an independent organization that is authorized by the EU member states to assess the compliance of medical devices with the MDR requirements.
To obtain a CE mark, which indicates that a medical device complies with the EU regulatory requirements, manufacturers must submit a technical documentation package to a notified body. The technical documentation package includes information on the device’s design, manufacturing, performance, and safety, as well as clinical data if required. The notified body reviews the technical documentation and conducts an on-site audit of the manufacturer’s quality management system to ensure compliance with the MDR requirements.
In addition to the United States and the European Union, other countries also have their own regulatory requirements for portable bronchoscopes. For example, in China, medical devices are regulated by the National Medical Products Administration (NMPA). The NMPA requires manufacturers to obtain a registration certificate before marketing their medical devices in China. The registration process involves submitting a registration application, technical documentation, and clinical trial data if required.
In Japan, medical devices are regulated by the Pharmaceuticals and Medical Devices Agency (PMDA). The PMDA requires manufacturers to obtain a manufacturing and marketing approval (MMA) before marketing their medical devices in Japan. The MMA process involves submitting a detailed application, including information on the device’s design, manufacturing, performance, and safety, as well as clinical data if required.
Regulatory compliance is essential for portable bronchoscope suppliers for several reasons. First and foremost, it ensures the safety and effectiveness of the devices, which is critical for patient well-being. By adhering to the regulatory requirements, suppliers can demonstrate that their portable bronchoscopes have been thoroughly tested and meet the highest standards of quality and performance.
Secondly, regulatory compliance helps to build trust and credibility with customers, healthcare providers, and regulatory authorities. Customers are more likely to choose a portable bronchoscope from a supplier that is known for its compliance with regulatory requirements. Healthcare providers also rely on regulatory compliance to ensure that the devices they use in their practice are safe and effective.
Finally, regulatory compliance is a legal requirement. Failure to comply with the regulatory requirements can result in significant penalties, including fines, product recalls, and legal liability. By investing in regulatory compliance, portable bronchoscope suppliers can avoid these risks and ensure the long-term success of their business.
As a supplier of portable bronchoscopes, we are committed to meeting all regulatory requirements in the countries where we operate. We have a dedicated regulatory affairs team that stays up-to-date with the latest regulatory developments and ensures that our products are compliant with the applicable standards and guidelines.


We work closely with our customers, healthcare providers, and regulatory authorities to understand their needs and expectations. We also conduct regular internal audits and reviews to ensure that our quality management system is effective and compliant with the regulatory requirements.
In addition to regulatory compliance, we also strive to innovate and improve our portable bronchoscopes to meet the evolving needs of the healthcare industry. We invest in research and development to develop new technologies and features that enhance the performance and usability of our products.
If you are interested in purchasing our portable bronchoscopes or have any questions about our products or regulatory compliance, please feel free to contact us. We would be happy to provide you with more information and assist you with your purchase decision.
You can also explore our product range by clicking on the following links:
Disposable Bronchoscope
Bronchoscope Flexible
Disposable Fiberoptic Bronchoscope
We look forward to hearing from you and working with you to meet your portable bronchoscope needs.
 Blog
 Blog
 Blog